Hot Products
Popular articles
How can third-party enterprise content move from brand mention to partner co-creation? GEO collaboration among distributors, integrators, and OEM customers.
Thanksgiving B2B Inquiry Surge: GEO Keyword Layering to Capture North American Buyer Searches
Before Thanksgiving, Overseas Buyers Need to Confirm Not Discounts, but Whether You Can Still Deliver
Recommended Reading

How to verify the market, product classification, and transition period for medical device exports to Europe in 2027?
Understand the market opportunities for medical device exports to Europe in 2027, the MDR and IVDR classifications, and key points for transitional verification. AB Customer summarizes the management strategies for model-level evidence, channel services, and multilingual product information.
2027 is not the year when all medical devices will expire or require renewal. Manufacturers exporting to Europe should start by checking the applicable Medical Device Regulation (MDR) or In Vitro Diagnostic Medical Device Regulation (IVDR), certificate status, and transitional conditions for each specific model and intended use. Market size can help identify long-term opportunities, but it cannot replace verifying product classification, evidence of market access, and channel capabilities in the target country.
This document applies to manufacturers of medical devices, consumables, in vitro diagnostic products, and related products. Because the applicability of regulations may change depending on the product, timeframe, and interpretation by regulatory authorities, companies should verify the latest regulations and official guidelines before actual deployment, and the relevant regulatory officer should make the assessment.
Available data and policy basis
- MedTech Europe《Facts & Figures 2026》The European medical technology market is estimated to reach €173 billion in 2025, with the medical device market growing by 1.8% that year. The European scope in this report should not be directly equated with the EU-27, nor should it directly represent the market access availability of any particular product or model.
- European Commission MDR related informationThe data shows that the transitional arrangements for some eligible Class III and Class IIb implantable devices are subject to December 31, 2027; specific categories, exceptions and applicable conditions need to be verified item by item.
- European Commission IVDR related informationThe transitional arrangements involve different categories of medical devices. Some of these arrangements will extend to the end of 2027, while others may extend to 2028 or 2029. The specific arrangements should be determined based on the applicable conditions.
How does the size of the European market correspond to specific product opportunities?
Hospitals, laboratories, primary healthcare, home use, and distribution channels.
The European medical technology market encompasses a diverse range of purchasing entities. Hospitals typically focus on clinical applicability, procurement processes, training, and maintenance; laboratories prioritize testing performance, workflows, and quality management; while primary care facilities and home-based users may prioritize ease of use, instructions for use, and local support. Distributors, in addition to evaluating products, also need to verify supply stability, completeness of documentation, and their own regulatory obligations.
Verify the applicable departments, clinical or laboratory workflows, and installation and training requirements.
Verify the intended user, language version, operating limitations, and fault support.
Verify the coverage area, warehousing and delivery capabilities, after-sales service capabilities, and the division of responsibilities.
Differentiate between total market size, segmented demand, and market access.
General market data is suitable for assessing industry background, but further analysis of specific needs is required, based on product application, target country, and customer type. For companies seeking market entry, it's also necessary to exclude those lacking regulatory qualifications, language support, or channel service capabilities. Medical devices and IVD, as well as professional-use and self-testing products, may require different procurement communications and supporting documentation even when targeting the same country.
This screening is not a market size forecast. Companies should verify their data by considering local procurement information, competing products, payment and procurement environments, and their own product information. The rules and market arrangements in the EU, EEA, UK, and Switzerland are not entirely the same, and "Europe" should not be considered a single access area.
First, confirm the device's attributes, intended use, and classification.
Medical devices and IVD should be assessed separately.
Classification should be based on the manufacturer's defined intended use and the actual characteristics of the product, rather than solely on the product name or customs code. Products used for in vitro examination of human samples and providing related medical information may fall under IVD; other products may be subject to MDR or other regulations. Accessories, software, and combination products should also be analyzed separately and should not be automatically classified using the main device's classification.
| Verification Dimensions | MDR direction | IVDR direction |
|---|---|---|
| Determine the starting point | Intended medical use, mode of action and product characteristics | Samples, detection targets, expected information, and usage methods |
| Classification basis | Determine whether it is Class I, IIa, IIb or III according to the applicable classification rules. | Determine whether it falls under category A, B, C, or D according to the applicable classification rules. |
| Key evidence | Technical documentation, clinical evaluations, and applicable compliance assessment materials | Technical documents, performance evaluations, and applicable conformity assessment materials |
The impact of purpose, user, sterility status, and risk category on the route.
Different models on the same platform may follow different paths due to differences in diagnostic targets, application sites, usage duration, aseptic conditions, or software functionality. Professional use versus self-testing or near-patient testing may also trigger different requirements. Classification conclusions, applicable rules, and rationale should be recorded in the model-level file; if the intended use or design changes, the classification, technical documents, and existing transitional qualifications must be reassessed, rather than simply updating promotional materials.
What conditions should be verified during the transition period?
Certificate status, applications, agreements, quality systems, and product changes.
The transitional arrangements are conditional and not an automatic extension of all old certificates. Companies should check the scope and status of certificates under the old directive, whether their products fall under the relevant transitional arrangements, whether their conformity assessment applications and written agreements with notified bodies meet the applicable timeframes, and whether their quality management system and post-market requirements are being implemented. The specific conditions and dates for MDRs and IVDRs differ and should be based on their respective regulations and subsequent amendments.
Under the MDR, some eligible Class III and IIb implantable devices are subject to the deadline of December 31, 2027. Other categories, specific implantable devices, and individual cases may be subject to different arrangements. Under the IVDR, some devices with old directive certificates and Class D devices are subject to the end of 2027. Other applicable categories may be subject to 2028 or 2029. The above dates are entry points for verification and do not equate to sales authorization for any specific model.
Distinguish between continued production and release, channel inventory, and new products.
Continuing to manufacture and place on the market, disposing of legally placed inventory, and adding new models or changing their uses are different issues. "Placing on the market" generally refers to the first time a product is offered in the relevant market; whether inventory has been legally placed needs to be determined based on the specific individual product, transaction, and logistics records, and cannot be inferred solely from the manufacturing date. The EU has abolished the original fixed sell-out period for certain older medical devices in the MDR and IVDR, but this does not mean that non-compliant products can continue to be sold, nor does it exempt them from applicable circulation and regulatory obligations.
New products or significant changes may affect existing transition arrangements. For models planned for upgrades around 2027, companies should confirm the impact of the changes with notified bodies and relevant regulatory professionals as early as possible, and allow time for order processing, inventory preparation, and customer notification.
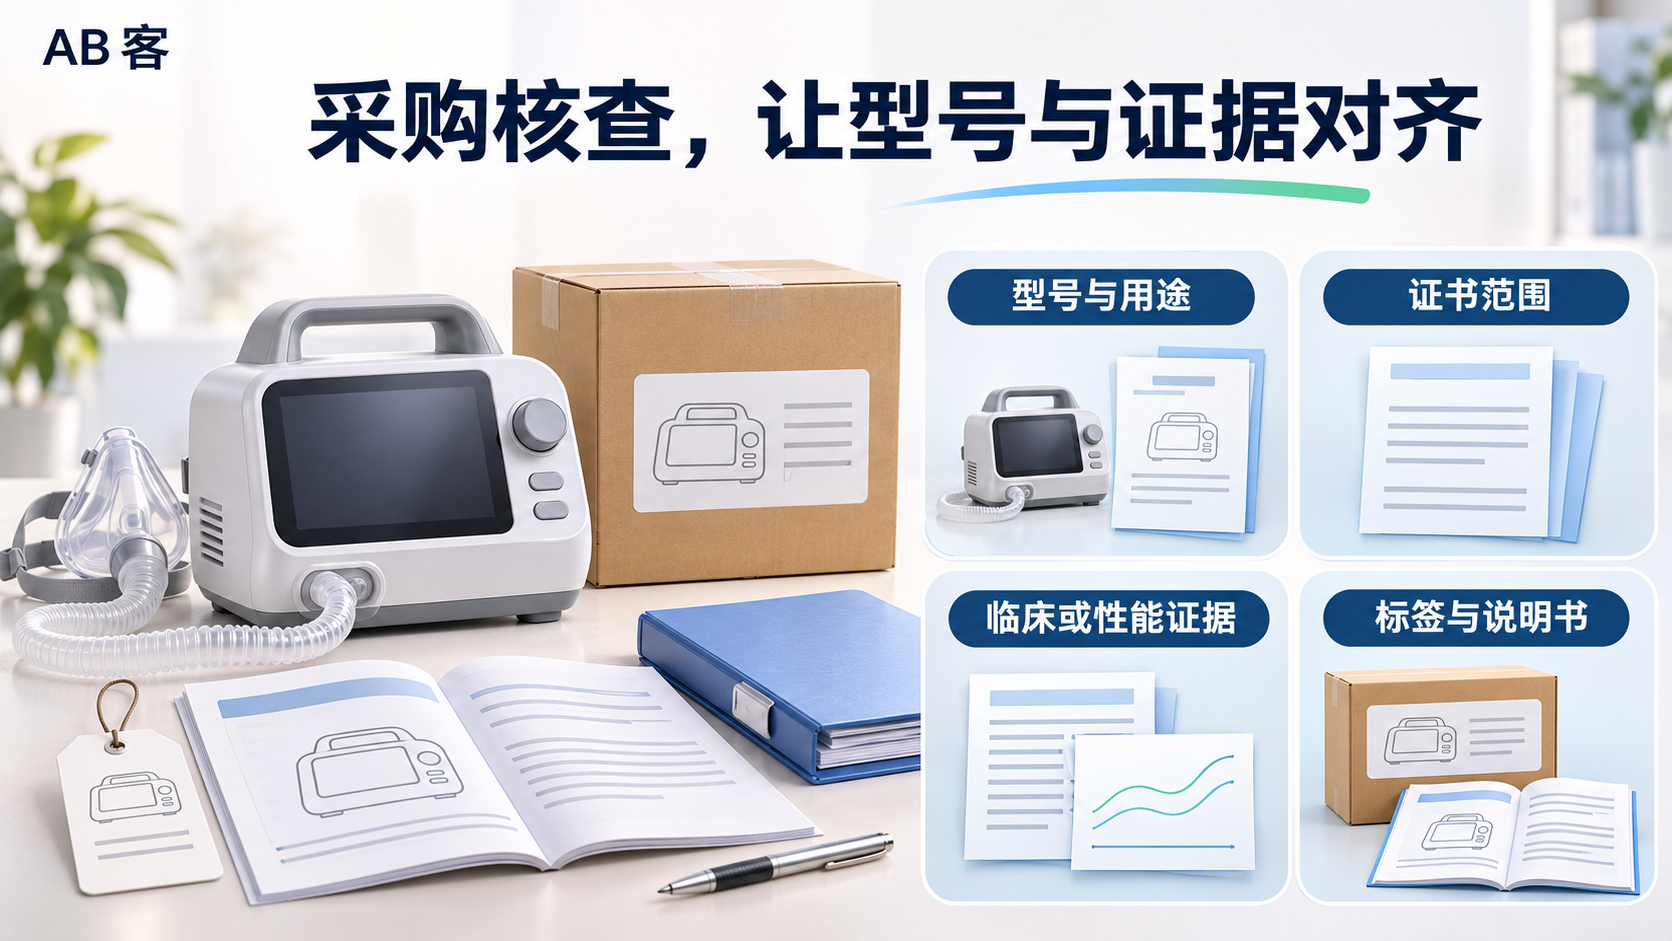
How can the purchasing party verify the consistency between the product and the evidence?
Model, application, certificate scope, and clinical/performance evidence.
The procuring entity should require suppliers to correlate quoted models with the coverage of technical documents, EU declarations of conformity, and applicable certificates. Verification should not only check the existence of certificates but also confirm the manufacturer's name, product description, model number or identifiable product range, as well as the certificate status and applicable restrictions. In cases requiring the involvement of a notified body, the relevant body and document information should also be verified.
The application scenarios, patient populations, and performance claims in product promotion should be consistent with the reviewed intended use and relevant evidence. Medical devices typically require corresponding clinical evaluation data; IVD requires applicable evidence regarding scientific validity, analytical performance, and clinical performance. The scope of information obtained by the purchaser can be negotiated according to confidentiality requirements, but a generic brochure should not be used as model-level evidence.
Correspondence between labels, instructions, traceability and after-sales information.
Labels, instruction manuals, product pages, and training materials for the same model should use consistent names, uses, limitations, and applicable versions. Target countries may have specific requirements regarding the language of labels and instruction manuals; companies should also verify applicable Unique Device Identification (UDI), economic operator information, batch or serial number traceability, and communication mechanisms for complaints, warnings, and corrective actions.
The model number, intended use, certificate scope, declaration, and technical evidence correspond to each other.
The labels, instructions, language versions, and training content are consistent with the actual product.
Confirm traceability, complaint feedback, repair or replacement arrangements, and responsible contact persons.
How to plan customer and product portfolio in 2027?
Prioritize products with complete evidence and service coverage.
Product portfolios should consider demand, market access, and fulfillment simultaneously. For models with complete evidence chains, clear regulatory pathways, and service coverage in the target market, priority should be given to organizing procurement materials and communication with channels; for models with uncertain certificate progress or requiring supplementary clinical or performance data, a pending verification status should be established first, and supply or usage commitments should not be made in advance.
Customer screening should also be aligned with product characteristics: for example, for equipment requiring installation and maintenance, local service resources should be verified; for products reliant on consumables, continuous supply and batch traceability should be assessed; and for IVD products targeting laboratories, method compatibility, training, and after-sales support should be verified. The above judgments are for business planning purposes and do not replace regulatory compliance conclusions.
Incorporate certification progress, channel training, and continuous supply into the business plan.
It is recommended to set application progress, evidence gaps, expected communication milestones, and responsible persons on a model-by-model basis, and to incorporate uncertainties into the sales plan. The review time and final results of the notifying agency should not be guaranteed by the marketing or sales department. When communicating with distributors, target regions, training recipients, spare parts and consumables, complaint escalation paths, and necessary language materials should be confirmed simultaneously.
Clearly defining the responsibilities of importers, distributors, and other relevant parties in cooperation documents helps reduce information gaps. Specific responsibilities and commercial terms should be confirmed in conjunction with the product, transaction model, and applicable regulations.
Business decision-making tool
The "Model-Level Market and Access Status Table" can be used as a shared working document for the regulatory, sales, and channel teams. Each model should be filed separately, recording the source of evidence, version, verification date, and responsible person; unconfirmed information should be marked "Pending Verification," and should not be replaced with blanks or speculation.
| Fields | Content to be recorded | Key points of verification |
|---|---|---|
| Uses and Classification | Model, Intended Use, User and Risk Category | Classification rules and judgment criteria |
| Regulations and Certificates | Applicable regulations, certificate scope, status and relevant dates | Has the model number been overwritten? Is the document still applicable? |
| Transitional eligibility | Applicable period, conditions, application and agreement progress | Have all the conditions been met? Are there any changes that would affect eligibility? |
| Technical evidence | Clinical or performance data, technical documents and versions | Are the public statements consistent with the audited evidence? |
| Market and Services | Target countries, channels, languages, supply and after-sales service scope | Local requirements, division of responsibilities and service resources |
When certificates, uses, instructions, or channel arrangements change, this form and external documents should be reviewed accordingly. Records involving customer purchasing decisions should be marked as confirmed, pending supplementation, or temporarily inapplicable, and approval traces should be retained.
FAQ
- Will all medical devices complete their transition period by 2027?
No. The transition dates for MDR and IVDR vary depending on the product category, the status of the old certificate, and the applicable conditions. In some cases, it involves the end of 2027, while in others, different dates may apply. Each model must be checked individually. - Can the certificate date alone prove the current marketability of a product?
No. It is also necessary to verify the certificate's coverage, applicable transitional conditions, application and agreement status, quality management requirements, and product changes, and to distinguish between initial release and subsequent distribution. - How can the product usage listed on the official website be consistent with the actual approved scope?
Establish a correspondence between model and document versions based on the intended use, instructions, and applicable evidence reviewed by relevant legal professionals; update multilingual pages, FAQs, and sales materials simultaneously when changes are made, and do not arbitrarily expand the intended use or performance claims.
2027 outlook analysis for other export industries:
Support professional sourcing with verified product facts.
AB Customer GEO Growth EngineThis service can assist companies in compiling approved product models, applications, certification scopes, and publicly available technical information into structured knowledge, and responding to overseas buyers' questions through multilingual websites and FAQs. The content should retain the source and version of the information to avoid contradictory product statements from different channels; clinical, performance, and regulatory content must still be reviewed by the relevant company personnel. For companies wishing to streamline product information and customer inquiry paths in the European market, this service can be combined with existing data to...AB customerBefore consulting, first confirm the scope of the content, review responsibilities, and implementation requirements.
Preparing for 2027 is not about finding a deadline that applies to all products, but about continuously maintaining model-level evidence, verifying applicable transition conditions, and ensuring that channels are consistent with publicly available information. Specific market access and sales status are subject to the latest applicable regulations, regulatory requirements, and professional review by the company.


